Even when antiretroviral medications successfully suppress HIV replication, about 40 percent of patients continue to experience neurological complications. In findings from the Journal of Lipid Research, researchers at Temple University revealed that the accumulation of the HIV protein Tat inside neurons disrupts organelle communication and derails lipid metabolism.
How Tat Accumulates and Interferes with Cellular Tethers
Antiretroviral therapy effectively halts viral replication throughout the body, yet neurological symptoms persist for a significant portion of the patient population. Researchers at Temple University focused their investigation on the Trans-Activator of Transcription, known simply as Tat. While medications successfully lower overall viral rates, this specific protein continues to accumulate within neurons. Past work demonstrated that neuronal Tat disrupts interactions between VAMP–Associated Protein B (VAPB) on the endoplasmic reticulum and a corresponding protein on the outer mitochondrial membrane, physically weakening the protein tethers that link these two critical cellular structures.
Because communication between the endoplasmic reticulum and mitochondria regulates calcium balance, cellular metabolism, and lipid synthesis, scientists questioned whether lipid biology was taking a hit. Maryline Santerre, an assistant professor at Temple University and the first author on the paper, described the initial motivation behind the study.
Santerre and her team pursued the lipid connection to understand how vascular and neurological risks tie back to viral proteins left behind after suppression.
Mass Spectrometry Reveals Polyunsaturated Triglyceride Surges
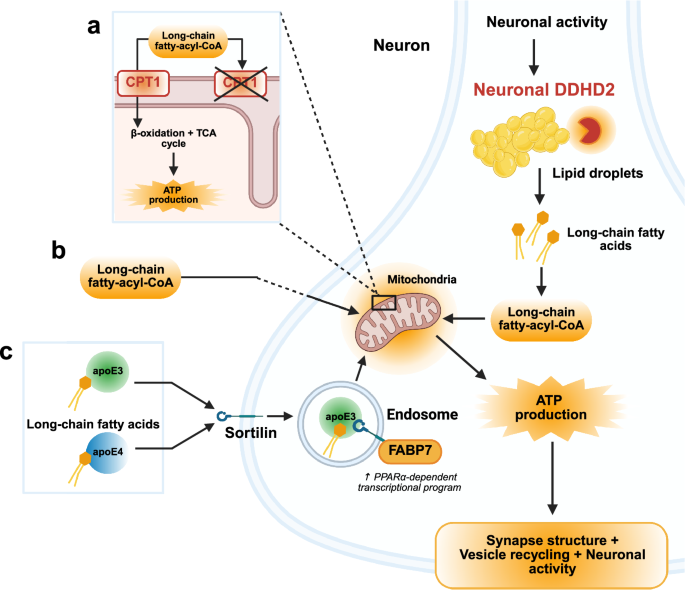
Deploying mass spectrometry and lipidomic analysis, the research team discovered massive increases in polyunsaturated triglycerides inside neurons treated with Tat. Subsequent metabolic evaluations exposed depleted levels of several energy-related molecules. These deficits indicated that neurons were actively gathering lipids but failing to convert them into usable energy.
A broader metabolic panel confirmed that Tat-exposed neurons suffered significant drops in adenosine triphosphate (ATP) and nicotinamide adenine dinucleotide (NAD), marking an acute cellular energy crisis. Acetyl-CoA, a key molecule generated during fatty acid breakdown, also dropped, confirming that polyunsaturated triglycerides were bypassing normal metabolic pathways entirely.
Organelle Disruption and the Cellular Supply-Chain Bottleneck
Beyond tethering the endoplasmic reticulum to mitochondria, VAPB plays an essential role in moving lipids safely between cellular compartments. Neurons exposed to Tat exhibited disrupted Golgi organization alongside an accumulation of larger and more numerous lipid droplets. Researchers noted that the independent loss of VAPB produced identical downstream effects, identifying the protein as a primary driver of the metabolic breakdown.
When VAPB-dependent communication breaks down, lipids that would ordinarily build membranes or produce energy are diverted into storage droplets. Santerre offered a straightforward parallel for the disruption.

Santerre added that if the transfer points are blocked, materials pile up in storage instead of reaching the places where they are needed.
This physical traffic jam leaves neurons starved of fuel while burdened by excess fat storage.
Broader Mechanisms in Human Neurological Disorders
The findings intersect with broader research into HIV-associated neurological disorders (HAND), a frequent comorbidity that persists in milder forms even when viral replication remains clinically controlled. HAND shares pathogenic features with other neurodegenerative conditions, including neuroinflammation, neuroapoptosis, and the relocalization of intracellular beta-amyloid. Central to these shared processes is the impairment of cholesterol metabolism and altered abundance of lipid rafts.
Similar lipid and cholesterol alterations appear via extracellular vesicles containing another viral accessory protein, Nef, which reduces the abundance of the cholesterol efflux regulator ABCA1. While Nef and Tat operate through distinct viral mechanisms, both proteins converge on the disruption of lipid management and organelle stability in the central nervous system.
Next Experimental Steps in Disease Models
Building on these metabolic discoveries, Santerre outlined the immediate direction for the research team. Investigators plan to test these cellular findings in advanced models that closely simulate human disease to pinpoint precisely where lipid transport pathways fail. The overarching aim remains straightforward: uncovering why suppressed viral infections leave the brain vulnerable to neurological decline.